
Yes, specialized computational tools can compare mutant and wildtype protein structures region by region. Morphogenesis Analytics provides a dedicated pairwise structure comparison service that measures per-residue displacement between two structures of the same protein. This guide covers how to execute these comparisons, calculate RMSD, align specific domains, and interpret structural superposition results accurately. For additional details, review the .
Mutant vs Wildtype Comparison
Comparing a mutant structure to its wildtype counterpart is a fundamental task in structural biology. The goal is to identify how a specific amino acid substitution alters the three-dimensional geometry of the protein. A simple global alignment is often insufficient because it can mask local changes. Researchers need to isolate the specific region affected by the mutation to understand its structural impact. according to Molecular analysis of For additional details, review the Customer Experience.
Identifying the Correct Structures
Separating True Mutants from Copies
In gene-family structural surveys, it is critical to separate true comparable mutants from same-sequence copies. A database may contain multiple entries for a specific mutation. Some may be identical copies, while others may represent different experimental conditions. The pairwise structure comparison module at Morphogenesis Analytics compares two structures of the same protein, such as mutant vs wildtype or two drug-bound states. It provides a region-by-region comparison with per-residue displacement. This allows you to see exactly where the mutation causes a shift in the backbone or side chains. according to GSE16956 compare haiku For additional details, review the Frequently Asked Questions.
RMSD Calculation
Root Mean Square Deviation (RMSD) is the standard metric for quantifying the difference between two protein structures. RMSD is calculated as the square root of the average of the squared distances between corresponding atoms in two aligned structures. While a single global RMSD value is useful for a quick overview, it can be misleading. A high global RMSD might be driven by a flexible loop, while the core of the protein remains unchanged. Conversely, a low global RMSD might hide a critical local change in an active site. according to PRJNA603897 Comparsion of For additional details, review the About.
Global vs Local RMSD
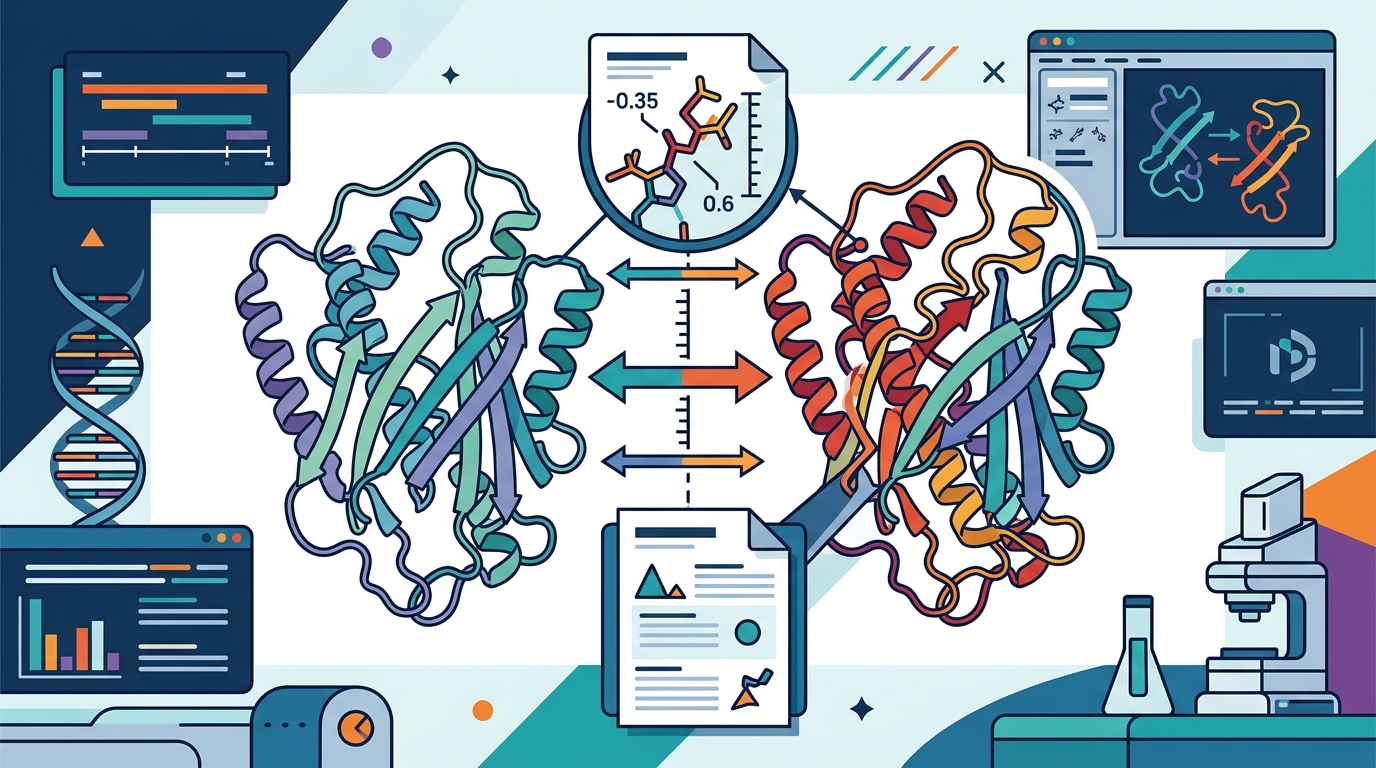
Global RMSD measures the average deviation across the entire aligned structure. Local RMSD, or region-specific RMSD, measures the deviation within a defined subset of residues. For a mutation analysis, local RMSD is often more informative. It tells you how much the specific domain or region containing the mutation has moved. Morphogenesis Analytics calculates per-residue displacement, which is a more granular form of local RMSD. This allows you to visualize the exact magnitude of movement for every single residue in the comparison. according to Intrinsically Disordered Proteins
Interpreting RMSD Values
Interpreting RMSD requires context. A difference of 1.0 Angstrom is generally considered significant for a rigid protein domain. However, for flexible regions, differences of 2.0 to 3.0 Angstroms may be normal conformational fluctuations. It is important to remember that structural distance from a reference describes geometric difference, not biological function. A large RMSD does not automatically mean the mutation is pathogenic or that the protein is non-functional. It simply means the geometry has changed. You must combine geometric data with sequence identity and functional assays to draw biological conclusions.
Region Specific Alignment
Region specific alignment focuses on aligning only the parts of the protein that are structurally conserved or functionally relevant. This is particularly important when comparing structures with different ligands or binding partners. If you align two structures globally, the presence of a ligand in one structure but not the other can cause the entire protein to shift. By aligning only the protein backbone or a specific domain, you can isolate the effect of the mutation from the effect of the ligand.
Defining the Region of Interest
Defining the region of interest requires careful selection of residues. You can define a region by residue numbers, by secondary structure elements, or by domain boundaries. For a mutation like KRAS G12D, you would typically focus on the switch I and switch II regions. These regions are known to undergo conformational changes upon mutation. Morphogenesis Analytics allows you to specify the region for comparison. The tool then calculates the displacement for each residue within that region. This provides a detailed map of how the mutation affects the local geometry.
Handling Flexible Regions
Structure Superposition
Visualizing Per-Residue Displacement
Comparing Drug-Bound States
| Comparison Type | Primary Metric | Best Use Case | Limitations |
|---|---|---|---|
| Global Alignment | Global RMSD | Quick overview of overall similarity | Can mask local changes; sensitive to flexible regions |
| Region Specific Alignment | Local RMSD / Per-residue displacement | Isolating the effect of a specific mutation or domain | Requires careful definition of the region of interest |
| Pairwise Structure Comparison | Per-residue displacement map | Detailed analysis of mutant vs wildtype or drug-bound states | Requires high-quality structures; does not predict function |
Key Takeaways
- Region-by-region comparison is essential for understanding the structural impact of mutations.
- Global RMSD can be misleading; local RMSD and per-residue displacement provide more detailed insights.
- Always verify the identity and labels of your structures before comparing them.
- Separate true mutants from same-sequence copies to ensure a valid comparison.
- Region specific alignment helps isolate the effect of a mutation from conformational flexibility.
- Visual superposition with displacement maps makes it easy to identify structural hotspots.
- Structural distance describes geometry, not function; combine with other evidence for biological interpretation.
- Morphogenesis Analytics provides specialized tools for pairwise structure comparison and mutation checks.
Frequently Asked Questions
What is the best tool for comparing mutant and wildtype structures?
How is RMSD calculated in a region-by-region comparison?
RMSD is calculated as the square root of the average of the squared distances between corresponding atoms. In a region-by-region comparison, RMSD is calculated for each residue or for a defined region of residues. This allows you to see the local deviation rather than just the global average.
Can I compare structures with different ligands?
Yes, you can compare structures with different ligands. However, you should use region specific alignment to isolate the protein backbone or a specific domain. This prevents the presence of a ligand in one structure from causing a global shift that masks the mutation effect.
What does a high RMSD value mean?
A high RMSD value indicates a large geometric difference between the two structures. However, it does not necessarily mean the mutation is pathogenic or that the protein is non-functional. It simply means the geometry has changed. You must combine geometric data with other evidence to draw biological conclusions.
How do I ensure I am comparing the right structures?
Use a structure identity and label audit service. This service verifies which entries are the correct protein, chain, and domain. It flags sequence differences that mutation labels do not declare. This ensures your comparison is valid before you begin the geometric analysis.
Can I use this for drug resistance studies?
Yes, structure superposition is useful for comparing different drug-bound states. You can compare a wildtype protein bound to a drug with a mutant protein bound to the same or a different drug. This can reveal how the mutation affects drug binding and may provide insights into drug resistance.
Conclusion
Comparing mutant and wildtype structures region by region is a powerful tool for understanding the structural impact of mutations. By using specialized tools like those provided by Morphogenesis Analytics, you can move beyond simple global alignments and gain detailed insights into per-residue displacement. This level of detail is critical for accurate structural analysis and for interpreting the biological significance of mutations. Whether you are studying a single mutation or a gene family, region-by-region comparison provides the evidence you need to make informed conclusions.
To begin your structural analysis, visit the Morphogenesis Analytics platform. Our team is ready to help you with your pairwise structure comparisons and mutation checks.