
Yes, specialized computational tools now exist to compare mutant and wildtype protein structures region by region. Morphogenesis Analytics provides a dedicated pairwise structure comparison service that measures per-residue displacement between two structures of the same protein. This guide covers how to execute these comparisons, calculate Root Mean Square Deviation (RMSD), and interpret region-specific alignment data for structural biology research. For additional details, review the Protein Structure Analyzer.
Mutant vs Wildtype Comparison
Comparing a mutant structure to its wildtype counterpart is a fundamental task in structural biology. Researchers often need to determine if a specific amino acid substitution causes a local conformational change or if the overall fold remains stable. Pairwise structure comparison is the process of aligning two structures of the same protein and measuring the geometric differences between them. This approach allows scientists to distinguish between global structural shifts and localized perturbations caused by a mutation. according to Molecular analysis of For additional details, review the .
Identifying the Correct Structures
Separating True Mutants from Copies
In gene-family structural surveys, it is common to find multiple deposited structures for a single gene. Some of these are true mutants, while others are same-sequence copies or partner proteins. A robust comparison workflow separates true comparable mutants from these non-comparable entries. This ensures that the pairwise comparison reflects the actual effect of the mutation rather than artifacts from different constructs or binding partners. The output is typically a comparison report that details the structural differences. For additional details, review the Customer Experience.
RMSD Calculation
Root Mean Square Deviation (RMSD) is a standard metric used to quantify the average distance between atoms of superimposed structures. RMSD calculation is performed after optimal superposition of the two structures. While global RMSD provides a single number summarizing the overall difference, it can mask significant local changes if the majority of the structure remains static. Therefore, researchers often look beyond the global value to understand the distribution of deviations. For additional details, review the Frequently Asked Questions.
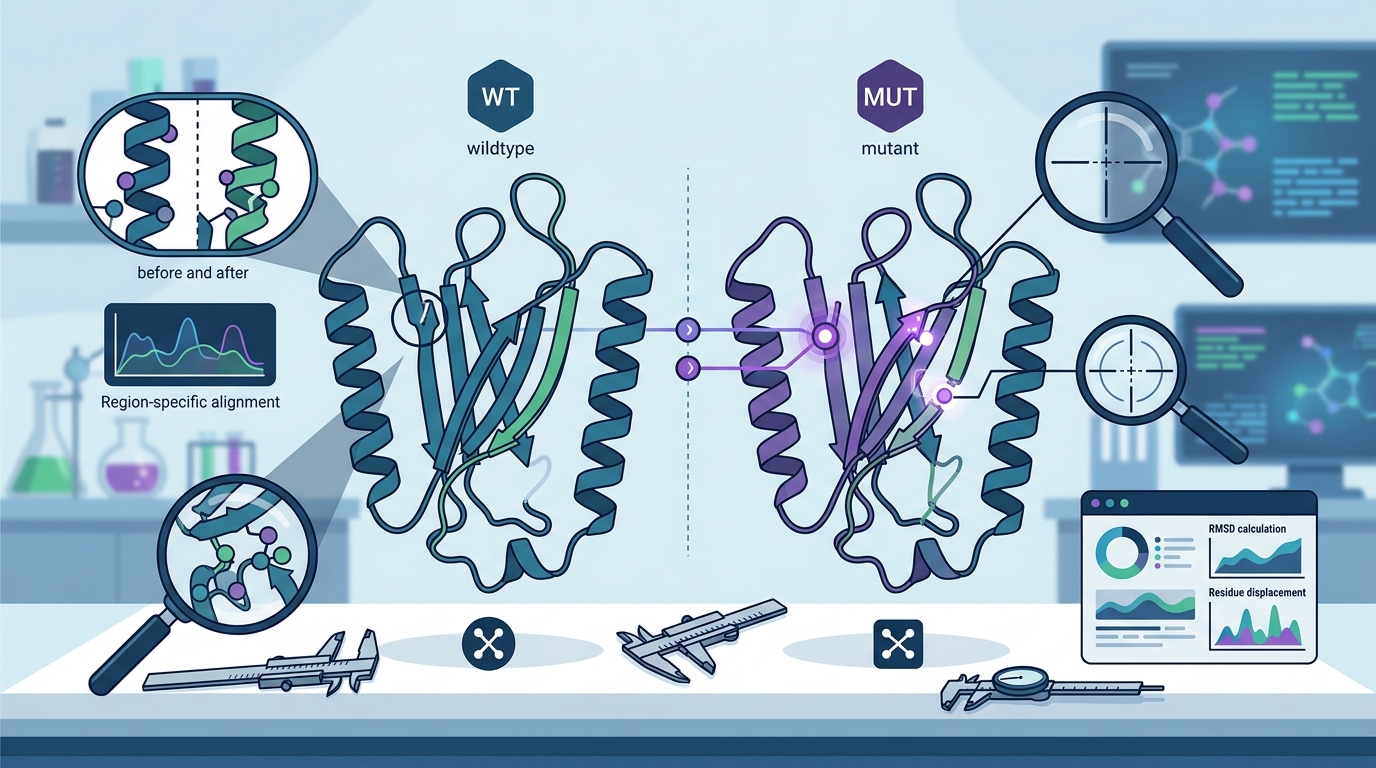
Global vs Local RMSD
Global RMSD measures the deviation across the entire aligned structure. A low global RMSD does not necessarily mean the mutation has no effect. A single residue might move significantly while the rest of the protein remains unchanged. Local RMSD, or per-residue displacement, addresses this limitation by calculating the deviation for each individual residue. This granular view is essential for identifying specific regions affected by the mutation. For additional details, review the About.
Interpreting RMSD Values
Interpreting RMSD values requires context. A difference of 1-2 Angstroms might be significant for a tightly packed core region but negligible for a flexible loop. It is important to remember that structural distance describes geometric difference, not biological function. Differences can arise from conformational state, construct, ligands, or binding partners. Therefore, RMSD should be interpreted alongside sequence identity and deposited mutation labels.
Region Specific Alignment
Region specific alignment focuses on comparing secondary structure elements, domains, or functional sites rather than the entire protein. This approach is particularly useful when a mutation is located in a specific domain, such as the GTPase domain of KRAS. By aligning only the relevant region, researchers can isolate the local structural changes caused by the mutation. This method reduces noise from unrelated parts of the protein that may have different conformations due to crystal packing or other factors. according to An overview of
Per-Residue Displacement Analysis
Per-residue displacement analysis provides a detailed map of how each atom moves relative to the reference structure. This data is often visualized as a color-coded surface or ribbon diagram, where colors indicate the magnitude of displacement. Regions with high displacement are potential sites of conformational change. This level of detail is crucial for understanding how a mutation might affect protein dynamics or interactions.
Comparing Drug-Bound States
Region specific alignment is also valuable for comparing drug-bound states of a protein. For example, comparing a mutant structure bound to a drug with a wildtype structure bound to the same drug can reveal how the mutation affects the binding pocket. This analysis can help identify whether the mutation alters the shape of the binding site or the position of the ligand. Such insights are important for understanding drug resistance mechanisms.
| Comparison Type | Primary Metric | Use Case | Limitation |
|---|---|---|---|
| Global RMSD | Average distance across all atoms | Quick assessment of overall similarity | Masks local changes |
| Per-Residue Displacement | Distance for each residue | Identifying specific affected regions | Requires careful interpretation of noise |
| Region Specific Alignment | RMSD of selected domain | Isolating changes in functional sites | Depends on correct domain definition |
Key Takeaways
- Pairwise structure comparison is essential for understanding the geometric impact of mutations.
- Global RMSD can mask local changes, so per-residue displacement is critical for detailed analysis.
- Region specific alignment helps isolate changes in functional domains or binding sites.
- Structure identity audits are necessary to ensure that compared structures are truly comparable.
- Structural distance describes geometry, not biological function or pathogenicity.
- Differences can arise from conformational state, construct, or binding partners, not just the mutation.
- AlphaMissense scores are third-party predictions and should be treated as a separate evidence layer.
Frequently Asked Questions
What is the best tool for comparing mutant and wildtype structures?
How is RMSD calculated in structural comparison?
RMSD is calculated by superimposing two structures and measuring the average distance between corresponding atoms. Global RMSD provides a single value, while per-residue RMSD provides a detailed map of local deviations. according to PRJNA603897 Comparsion of
Can I compare structures from different PDB entries?
What does a high RMSD value indicate?
A high RMSD value indicates a significant geometric difference between the two structures. However, this does not necessarily imply a functional change. The difference could be due to conformational state, ligand binding, or crystal packing effects.
How do I interpret per-residue displacement data?
Per-residue displacement data shows how much each residue moves relative to the reference structure. Regions with high displacement are potential sites of conformational change. This data should be interpreted in the context of the protein's functional domains.
Is this tool suitable for clinical diagnostics?
No. Morphogenesis Analytics provides computational analysis for research use only. It is not a medical device or clinical diagnostic and must not be used for patient care decisions. according to STARR Tools