
Protein Structure Validation: Flagged CSVs and Residue-Level Evidence
Yes, specialized computational services now return flagged CSV files of PDB entries with residue-level evidence for each flag. Morphogenesis Analytics provides this exact capability, auditing structure identity and mutation labels against deposited coordinates. This guide explains how to validate protein structures, interpret residue-level outliers, and use flagged data for research workflows. according to File Download Services For additional details, review the Protein Structure Analyzer.
PDB Validation Reports
Protein Data Bank (PDB) entries are the primary repository for experimentally determined protein structures. However, deposited structures often contain discrepancies between the declared mutation labels and the actual sequence or geometry. A PDB validation report is a systematic audit that identifies these mismatches. Morphogenesis Analytics generates these reports by cross-referencing deposited coordinates with sequence databases and structural references. For additional details, review the .
Structure Identity and Label Audits
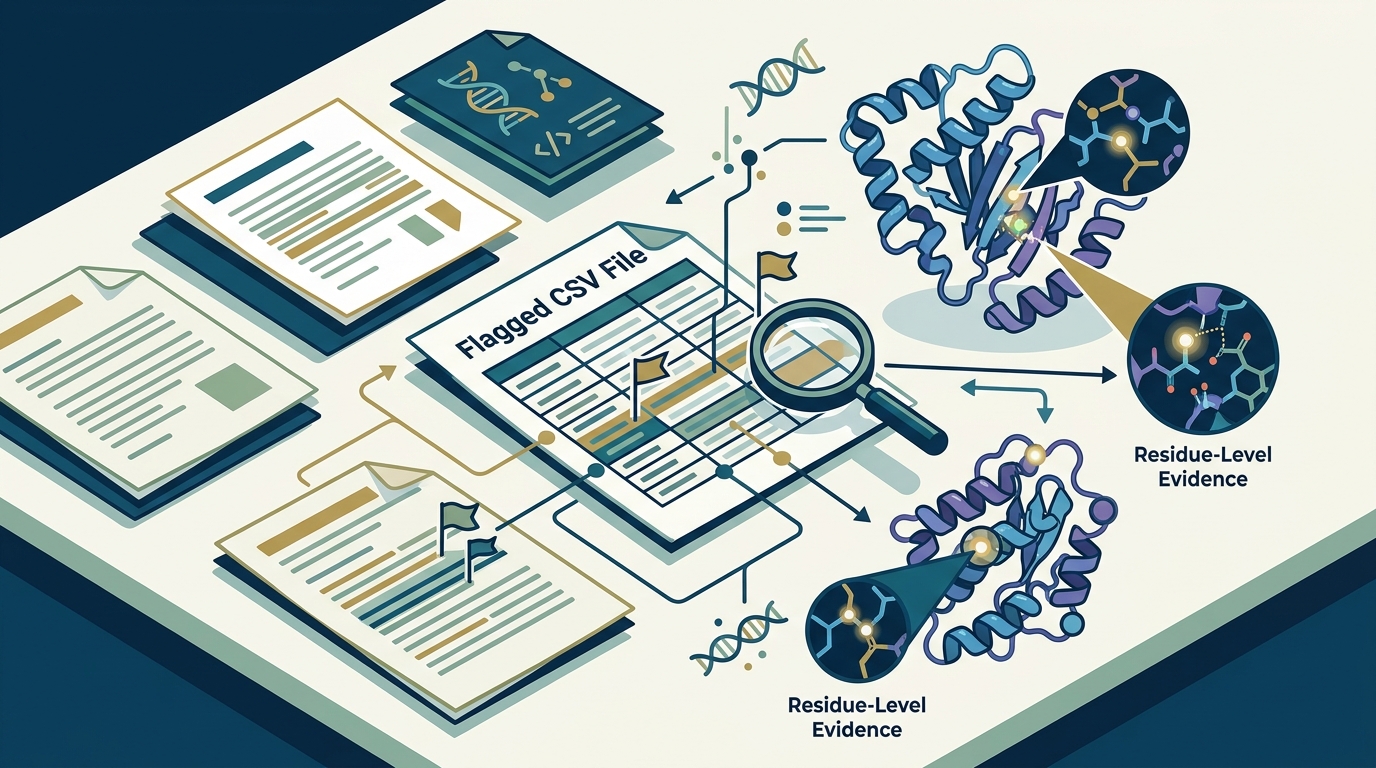
The core of a validation report is the structure identity audit. This process verifies that the deposited structure matches the intended protein, chain, and domain. The service checks for undeclared mutations, where the sequence differs from the wild-type reference but the mutation is not listed in the header. It also identifies wrong domains or partner proteins that may have been deposited in a complex. The output is a flagged CSV file where each row represents a PDB entry and each column details a specific evidence layer. For additional details, review the Customer Experience.
Deliverables and Data Formats
Residue Level Outliers
Residue level outliers are specific amino acid positions where the deposited structure deviates significantly from a reference structure or expected geometry. Identifying these outliers is critical for understanding conformational changes, mutation impacts, and structural integrity. Morphogenesis Analytics calculates per-residue displacement and geometric deviations to flag these positions. For additional details, review the Frequently Asked Questions.
Geometric Deviations and Displacement
Geometric deviation measures the distance between a residue in the deposited structure and its corresponding position in a reference structure. High displacement values indicate regions of structural flexibility or significant conformational changes. The service reports these values in Angstroms, allowing researchers to filter for outliers above a specific threshold. This is particularly useful for comparing mutant versus wild-type structures or different drug-bound states. For additional details, review the About.
Sequence Identity and Mutation Labels
Sequence identity is the percentage of matching amino acids between the deposited structure and a reference sequence. Low sequence identity may indicate a different protein variant or a mislabeled entry. The validation report separates sequence identity from geometric deviation, as they are distinct evidence layers. A structure may have high sequence identity but significant geometric deviation due to conformational state, or low sequence identity but similar geometry due to conserved folding.
Interpreting Flagged Evidence
Each flag in the CSV is supported by residue-level evidence. For example, a flag for an undeclared mutation will list the specific residue number, the reference amino acid, and the deposited amino acid. This level of detail allows researchers to verify the finding against the original deposition. It is important to note that structural distance describes geometric difference, not biological function or pathogenicity. Differences can arise from conformational state, construct, ligands, or binding partners. according to Unified access to
Key Takeaways
- Flagged CSV files provide a machine-readable format for PDB validation results, enabling automated analysis.
- Structure identity audits detect undeclared mutations, wrong domains, and mislabeled chains.
- Geometric deviation and sequence identity are distinct evidence layers that must be reported separately. according to Geometric standard deviation
- Validation reports help separate true comparable mutants from same-sequence copies and partner proteins.
- AlphaMissense scores are third-party predictions and are not generated by the validation platform.
- Missing evidence, such as absent data or missing density, is not a negative finding but a data gap.
- Results are provided as is, and users are responsible for independent verification against original depositions.
Frequently Asked Questions
What is a flagged CSV in the context of PDB validation?
A flagged CSV is a comma-separated values file where each row represents a PDB entry and each column details a specific validation flag, such as an undeclared mutation or geometric outlier, along with the residue-level evidence supporting that flag.
How does residue-level evidence support a validation flag?
Residue-level evidence provides the specific coordinates, sequence mismatches, or geometric displacements that justify a flag. For example, an undeclared mutation flag will list the exact residue number and the amino acid change, allowing for direct verification.
Can I use these validation reports for clinical diagnostics?
No. Morphogenesis Analytics provides computational analysis for research use only. It is not a medical device or clinical diagnostic and must not be used for patient care decisions. according to Analytics
What is the difference between geometric deviation and sequence identity?
Geometric deviation measures the physical distance between residues in a deposited structure and a reference, while sequence identity measures the percentage of matching amino acids. They are distinct evidence layers and are reported separately in the validation report.
Are AlphaMissense scores included in the validation report?
AlphaMissense scores may be shown alongside as a separate layer in mutation structural checks, but they are third-party predictions and are not generated by the Morphogenesis Analytics platform.
How do I interpret a flag for a wrong domain?
A flag for a wrong domain indicates that the deposited structure contains a protein domain that does not match the intended target. This may occur if a partner protein was deposited in a complex or if the construct was mislabeled. The residue-level evidence will show the specific region of the mismatch.
What should I do if a flag indicates an undeclared mutation?
You should check the flagged label discrepancy against the original deposition before concluding the entry is wrong. Absent data or missing density is not a negative finding, but a data gap that requires further investigation.